Your device has been on the European market for years under a Medical Device Directive certificate. The certificate’s printed expiry date has passed, your distributors are asking whether they can keep ordering, and someone in the business has heard that “the MDR deadline moved to 2028”. Whether that applies to your device depends on its class, on paperwork you should have completed in 2024, and on whether you have changed the device since.
This article sets out the MDR transition deadline for legacy devices exactly as Regulation (EU) 2023/607 wrote it into Article 120 of Regulation (EU) 2017/745 (the MDR), the conditions that keep a device inside the extension, and the mistakes that push manufacturers out of it.
What Regulation (EU) 2023/607 changed
Regulation (EU) 2023/607 was adopted on 15 March 2023, published in the Official Journal (L 80, p. 24) on 20 March 2023 and entered into force the same day. Its recitals are direct about the reason: the overall capacity of notified bodies was still not sufficient to carry out conformity assessment for the large number of devices holding Directive certificates, and the extension was meant to reduce the risk of shortages without lowering safety requirements.
It did three things to the MDR:
- Extended the validity of certificates issued under Directive 90/385/EEC (AIMDD) and Directive 93/42/EEC (MDD).
- Replaced the single transition date with risk-based dates in a new Article 120(3a) and (3b).
- Made the extension conditional on steps the manufacturer had to take by fixed dates in 2024 (Article 120(3c)).
It also removed the “sell-off” limit on devices already placed on the market (see below).
The MDR transition deadline by device class
Under Article 120(3a), devices with a valid MDD or AIMDD certificate may be placed on the market or put into service until:
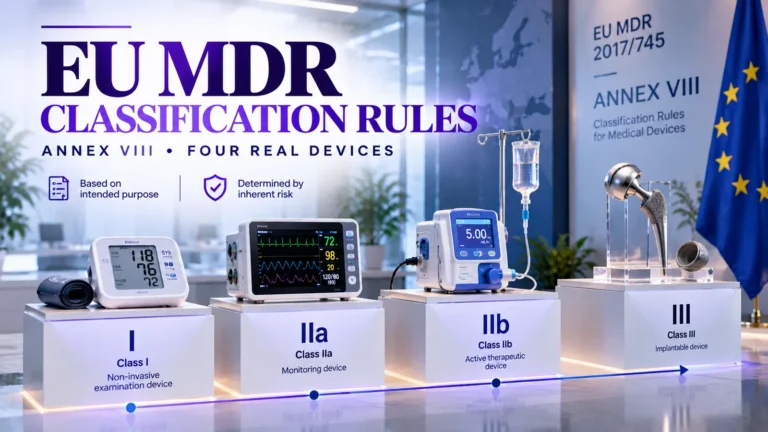
- 31 December 2027 — all class III devices, and class IIb implantable devices except sutures, staples, dental fillings, dental braces, tooth crowns, screws, wedges, plates, wires, pins, clips and connectors.
- 31 December 2028 — all other class IIb devices (including the implantable exceptions just listed), class IIa devices, and class I devices placed on the market in sterile condition or with a measuring function.
Article 120(3b) covers a different group: devices that needed no notified body under the MDD, whose declaration of conformity was drawn up before 26 May 2021, and which now need a notified body under the MDR (typically devices up-classified by the MDR rules). These may be placed on the market until 31 December 2028.
Note the wording: the date limits placing on the market. It is not a date by which you merely need to have applied. To keep supplying new units after it, you need an MDR certificate.
Which certificates were extended
Article 120(2) extends certificates issued under the AIMDD or MDD from 25 May 2017 that were still valid on 26 May 2021 and have not been withdrawn since. They remain valid beyond the date printed on them until the relevant date in Article 120(3a).
Certificates that had already expired before 20 March 2023 were rescued only if one of two things was true:
- Before the expiry date, the manufacturer and a notified body had signed a written agreement under Annex VII, Section 4.3, second subparagraph, for the device or a device intended to substitute it; or
- A competent authority had granted a derogation under Article 59(1) or required the conformity assessment under Article 97(1).
The conditions in Article 120(3c)
The extended dates apply only if all of these conditions are met:
- (a) the devices continue to comply with the MDD or AIMDD, as applicable;
- (b) there are no significant changes in the design and intended purpose;
- (c) the devices do not present an unacceptable risk to the health or safety of patients, users or other persons, or to other aspects of public health;
- (d) no later than 26 May 2024, the manufacturer had put in place a quality management system in accordance with Article 10(9) MDR;
- (e) no later than 26 May 2024, the manufacturer or authorised representative had lodged a formal application with a notified body under Annex VII, Section 4.3, first subparagraph, for the legacy device or a device intended to substitute it, and no later than 26 September 2024 had signed a written agreement with that notified body.
Points (d) and (e) are now historical facts. If they were not met by those dates, the extension does not apply to the device. Points (a) to (c) are continuing conditions: they must stay true every day until your MDR certificate is issued.
Surveillance, vigilance and EUDAMED for legacy devices
Legacy status does not mean Directive rules only. Under Article 120(3d), the MDR requirements on post-market surveillance, market surveillance, vigilance and registration of economic operators and devices apply in place of the corresponding Directive requirements.
Under Article 120(3e), the notified body that issued the Directive certificate remains responsible for appropriate surveillance unless the manufacturer agrees with an MDR-designated notified body that it will take over. No later than 26 September 2024, the notified body that signed the written agreement became responsible for surveillance of the devices it covers. That notified body is not responsible for conformity assessment activities carried out by the issuing notified body.
Regulation (EU) 2024/1860 added a further obligation that applies to legacy and MDR devices alike from 10 January 2025: Article 10a MDR. If you anticipate an interruption or discontinuation of supply of a device (other than a custom-made device) and it is reasonably foreseeable that this could result in serious harm or a risk of serious harm to patients or public health in one or more Member States, you must inform the competent authority where you or your authorised representative are established, and the economic operators, health institutions and healthcare professionals you supply directly, at least six months beforehand, other than in exceptional circumstances. This matters if your plan is to let some legacy products lapse rather than certify them.
Sell-off removed and custom-made implants
Before 2023/607, devices placed on the market under the Directives could only be made available until a fixed sell-off date. Article 120(4) now has no time limit: devices lawfully placed on the market under the Directives, or under Article 120(3), (3a), (3b) or (3f), may continue to be made available on the market or put into service. A unit placed on the market on 30 December 2027 can therefore still be sold by a distributor afterwards; a unit first placed on the market in 2028 under an expired extension cannot.
Article 120(3f) allowed class III custom-made implantable devices to be placed on the market without a notified body certificate until 26 May 2026, subject to the same 2024 application and agreement dates. That window has closed.
Where this goes wrong
- Treating the date as a submission date. 31 December 2027 or 2028 is the last day to place legacy units on the market, not the day your file is due. Notified body questions, clinical evidence gaps and audit findings all happen before it. Build the plan backwards from the date.
- Making a “small” change that is significant. Condition (b) is lost through design or intended-purpose changes. MDCG 2020-3 Rev.1 (May 2023) gives flowcharts for intended purpose, design, software, materials and sterilisation changes; a change is non-significant only if every applicable chart says so. Ask the notified body in writing when in doubt.
- An application that does not cover the product actually sold. Point (e) requires the application to cover the legacy device or a device intended to substitute it. Variants, accessories or kits missing from the application and agreement may not be covered.
- Running vigilance and PMS under the old Directive logic. Article 120(3d) applies MDR post-market surveillance and vigilance now. Notified bodies auditing surveillance will look for MDR-style PMS plans and reports.
- Assuming the class has not changed. The dates in Article 120(3a) follow the device’s class. Confirm it against the MDR rules early, because an up-classified device may need a different conformity route and more clinical data.
- Waiting for the proposal. The Commission’s simplification proposal of 16 December 2025 is not law. Plan to the dates in force today.
Frequently asked questions
What is the MDR transition deadline for class III devices?
Under Article 120(3a) MDR, as amended by Regulation (EU) 2023/607, class III devices with a valid MDD or AIMDD certificate may be placed on the market until 31 December 2027, provided all the conditions in Article 120(3c) are met. The same date applies to most class IIb implantable devices. After that date, new units need an MDR certificate.
Which devices have until 31 December 2028?
Class IIb devices other than the implantables with a 2027 date, class IIa devices, and class I devices that are sterile or have a measuring function, if they hold a valid Directive certificate. The 2028 date also applies under Article 120(3b) to devices self-declared under the MDD before 26 May 2021 that now need a notified body under the MDR.
My MDD certificate shows an expiry date that has passed. Is it still valid?
Possibly. Article 120(2) extends certificates issued from 25 May 2017 that were valid on 26 May 2021 and not withdrawn, up to the Article 120(3a) date. Certificates that expired before 20 March 2023 are covered only if a written agreement was signed before expiry or a competent authority acted under Article 59(1) or 97(1). All 120(3c) conditions must also be met.
Can distributors keep selling legacy stock after 2027 or 2028?
Yes, for units already placed on the market lawfully. Regulation (EU) 2023/607 removed the earlier sell-off limit, so Article 120(4) lets such devices continue to be made available or put into service without a time limit. What ends on the deadline is the manufacturer’s ability to place new legacy units on the market.
Will the proposed MDR revision change these deadlines?
The Commission published a proposal to simplify the MDR and IVDR on 16 December 2025. It is a proposal under negotiation, not adopted law, and nothing in it applies until the European Parliament and Council adopt a final text. Plan to the dates currently in Article 120 and track the legislative process separately.
If you are working towards a 2027 or 2028 date and want a second view of your plan, Medfins has supported more than 100 clients across 175+ regulatory projects and 250+ products. Our CE marking under EU MDR service covers gap assessment, technical documentation and notified body preparation for legacy devices; contact our regulatory team to discuss your portfolio.