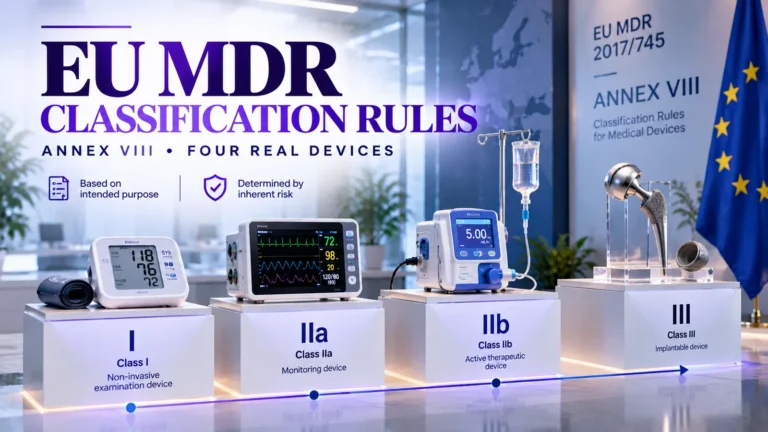
EU MDR Classification Rules: Annex VIII Applied to Four Real Devices
How to classify a medical device under Annex VIII of the EU MDR, with worked examples for needles, dressings, monitors and software, and the usual errors.
How to classify a medical device under Annex VIII of the EU MDR, with worked examples for needles, dressings, monitors and software, and the usual errors.
Which notified body should you apply to for CE marking? A step-by-step selection method using NANDO designation codes, Article 52 routes and fees.
A manufacturing or import licence does not let you sell onward. MD-42 wholesale registration does. Who needs it, what the application requires, and where it goes wrong.
BIS and CDSCO are not alternatives. Two authorities, two separate questions, and a product can need one, both or neither. How to work out which applies to yours.
What an MDSAP audit actually examines: the seven-process model, how nonconformities are graded numerically, realistic cost and timeline, and how to prepare.
FDA denied a 510(k) exemption for radiology CAD and CADt software, proposed eSTAR for PMAs and opened a robotics draft guidance. Three actions, two comment deadlines in November.
EUDAMED registration for devices already on the EU market closes on 28 November 2026. What MDR and IVDR manufacturers, authorised representatives and importers must check now.
MDSAP explained: the five participating authorities, why Canada makes it mandatory, why the EU does not accept it, the seven-process audit model, the 2026 changes, and how EN ISO 13485 differs from ISO 13485.
What an Indian Authorised Agent does under the Medical Devices Rules 2017, the MD-14 to MD-15 pathway, agent liability, and why appointing your distributor as agent can cost you the Indian market.